8th International Electronic Conference on Synthetic Organic Chemistry. ECSOC-8. 1-30 November 2004. http://www.lugo.usc.es/~qoseijas/ECSOC-8/
[A024]
Oxidative Iodination of Various Deactivated Arenes with I2 or KI in Concentrated Sulfuric Acid, and with Sodium Periodate Added as the Oxidant
Lukasz Kraszkiewicz, Maciej Sosnowski and Lech Skulski*
Chair and Laboratory of Organic Chemistry, Faculty of Pharmacy, Medical University,
1 Banacha Street, PL 02-097 Warsaw, Poland.
* To whom correspondence should be addressed; Tel./Fax: +(48) 22 5720643;
E-mail: [email protected]
Abstract: A number of more or less deactivated arenes, ArH, were effectively mono- or diiodinated with strongly electrophilic I+ reagents prepared from either diiodine or potassium iodide and with added sodium periodate, and reacted for 30 min at 25 – 30 °C in concd (min. 95%) sulfuric acid. The iodinating I+ solutions thus obtained were always applied in a 10% excess in respect to the reacted arenes, with using either the “direct” method or the “inverse” method of aromatic iodination explained in the text. The yields for the iodinated pure products varied from 17% to 91%.
Keywords: Oxidation, diiodine, sodium periodate as oxidant, oxidative iodination, deactivated arenes, iodoarenes
Introduction
Aromatic iodides, ArI, are generally more reactive, though also more costly, than the respective bromides and chlorides. There is a considerable number of different methods, direct and indirect, for their synthesis [1], and they are widely used in organic synthesis in chemical laboratories and, to a lesser extent, in chemical industry. Moreover, ArI are able to form a large variety of aromatic hypervalent iodine derivatives, which found quickly increasing applications in modern organic synthesis [2]. Our two latest reviews [3, 4] relate and explain a variety of aromatic oxidative iodination methods devised in our labolatory since 1990. See also Refs [5 – 8] for our newest papers on the oxidative iodination of various aromatics, published in 2004.
In our newest work [8], two “model” deactivated arenes, benzoic acid and nitrobenzene, were monoiodinated within one hour at 25 – 30 °C, with strongly electrophilic I+ (i.e. iodine(I) hydrogensulfate, IOSO3H) reagent, prior prepared from diiodine and various oxidants (CrO3, KMnO4, active MnO2, HIO3, NaIO3, or NaIO4) in 90% (v/v) concentrated sulfuric acid (ca. 75 mol% H2SO4). Next, only an I2/NaIO3/90% (v/v) conc. H2SO4 exemplary system was used to effectively mono- or diiodinate a considerable number of more or less deactivated arenes. All former papers dealing with the direct iodination of deactivated arenes were there briefly reviewed and commented. We had deduced that the following stoichiometries were obeyed in the iodinating reactions:
|
|
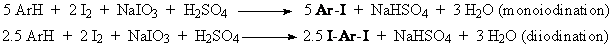
For nitrobenzene, benzoic acid and other deactivated arenes, which hardly can be diiodinated by a large excess of the iodinating solutions, a “direct” method of aromatic monoiodination can be the most successful for the reactions. All the monoiodinated arenes, taken in strictly stoichiometric amounts (0% excess), were always reacted with the iodinating solutions used in the excesses: 10, 50, or 100% - the last percentage was applied only for nitrobenzene (a “model” strongly deactivated arene). Using the “direct” method of aromatic iodination, benzoic acid or nitrobenzene, etc. were added at once to the appropriate, prior prepared, iodinating solutions, and the stiring was continued for one hour at 25 – 30 °C. The reactions were quenched by pouring the final reaction mixtures into ice-water. The precipitated crude products were collected by filtration, washed well with cold water, air-dried, and recrystallized from organic solvents to give the monoiodinated arenes in good or excellent yields [8]. However, for some weakly deactivated arenes, forming readily the hardly separable mixtures of mono- and diiodinated products, e.g. for benzoic acids or nitrobenzenes para-substituted with the methoxy or methyl group, an “inverse” method of their monoiodination was definitely preferable. The reacted weakly deactivated arenes, ArH, were suspended in definite volumes of 90% (v/v) concd H2SO4, kept at 25 – 30 °C with stirring, and appropriate iodinating solutions were very slowly added dropwise within 45 minutes, and the stirring was continued at the same temperature for a further 15 minutes. The reactions were quenched by pouring into ice-water, and the following workups and purification were the same as above. We also established experimentally that for the most effective diiodination reactions of some moderately deactivated arenes, the use of the aforementioned “direct” method was preferable; see Table 2 in Ref. 8. The reacted arenes were added at once to appropriate iodinating solutions, and the stirring was continued for two hours at 25 – 30 °C. The reactions were quenched by pouring into ice-water, and the following workups and the purification were the same as above. Only few iodinating reactions had to be carried out at 0 – 5 °C, for the reasons explained in Ref. 8. In this way, all the purified mono- or diiodinated deactivated arenes were previously obtained in 33 – 85% yields [8].
In order to extend the scope of our former oxidative iodination procedures [8], in the present work we have carried out quite numerous oxidative iodination reactions with a large variety of more or less deactivated arenes (Tables 1 – 3), with using either I2/NaIO4/concd (min. 95%) H2SO4 or KI/ NaIO4/concd (min. 95%) H2SO4 iodinating liquid systems. We have applied, when necessary, either the “direct” or the “inverse” iodination methods explained above and in Ref. 8. So far, the iodinating I+ solutions have always been applied in this work only in a 10% excess, in respect to the reacted arenes taken in strictly stoichiometric amounts (0% excess). We are planning to further improve our present iodinating experiments, before their final publishing.
Results and Discussion
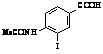
A number of deactivated arenes, ArH, shown in the Table 1, can hardly be diiodinated, even by large excesses of iodinating solutions. Hence, they were ”directly” monoiodinated either with I2/NaIO4/concd (min. 95%) H2SO4 system or with KI/NaIO4/concd (min. 95%) H2SO4 system to give, after workups and purification (see experimental part), the purified ArI in 43-91% yields. It is remarkable that the both iodinating systems, always with using the iodinating solutions containing I+ intermediates in a 10% excess, afforded nearly the same final yields. The following stoichiometries were obeyed for the monoiodination reactions:
|
|
|
|
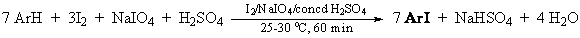
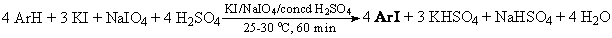
When the iodination reactions were complete, they were quenched by pouring into ice-water. However, the monoiodination of benzaldehyde should be carried out in 90% (v/v) concd H2SO4, under the same remaining reaction conditions, since the analogous reaction carried out in concd (min. 95%) H2SO4 afforded 3-IC6H4CHO notably admixed by 3-IC6H4COOH (TLC, NMR). It is due to the fact that the warm NaIO4/concd (min. 95%) H2SO4 system is apparently a stronger oxidant than the corresponding NaIO4/90% (v/v) concd H2SO4 system.
Table 1. Monoiodinated pure products prepared
|
ArH |
ArI |
Yield (%) |
Mp (oC) Solvent |
Lit, mp (oC) |
|
|
I2/NaIO4 |
KI/NaIO4 |
||||
|
|
|
80 |
82 |
186-188 CCl4 |
185-188 |
|
|
|
91 |
86 |
243-244 aq. AcOH |
243-245 |
|
|
|
82 |
84 |
214-215 aq. AcOH |
215-217 |
|
|
|
67 |
65 |
267-268 EtOH |
258-259 265-266 |
|
|
|
35 |
37 |
228-230 aq. EtOH |
230 |
|
|
|
45 |
43 |
51-52 petr. ether |
50-52 |
|
|
|
69 |
70 |
186-187 aq. EtOH |
185-187 |
|
|
|
72 |
71 |
152-153 H2O |
151-153 |
|
|
|
42a |
43a |
54-55 petr. ether |
53-56 |
|
|
|
54 |
56 |
58-60 petr. ether |
not found |
|
|
|
61 |
63 |
113-116 EtOH |
114-117 |
|
|
|
48 |
50 |
35-36 petr. ether |
35-38 |


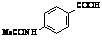


a) The reaction should be carried out with using 90% (v/v) concd H2SO4, under the same remaining reaction conditions.
Deactivated arenes, ArH, shown in the Table 2 should preferably be monoiodinated only by the ”inverse” method; when the ”direct” method was applied, the crude products thus obtained were notably contaminated by diiodinated side products, and the following repeated recrystallizations considerably lowered the yields of the desirable ArI. All the arenes from Table 2 were reacted, as above, only with the iodinating solutions containing I+ intermediates in a 10% excess. The final yields of ArI thus obtained varied from 56 up to 82%.
Table 2. Monoiodinated pure products prepared
|
ArH |
ArI |
Yield (%) |
Mp (oC) Solvent |
Lit, mp (oC) |
|
|
I2/NaIO4 |
KI/NaIO4 |
||||
|
|
|
79 |
78 |
210-212 EtOH |
208-212 |
|
|
|
80 |
81 |
247-249 EtOH |
233-234 242-243 |
|
|
|
65a |
67a |
96-98 EtOH |
95-98 |
|
|
|
56 |
57 |
112-113 EtOH |
112-114 |
|
|
|
72 |
70 |
53-55 EtOH |
54-56 |
|
|
|
81 |
82 |
98-99 EtOH |
97-99 |
|
|
|
67 |
66 |
92-93 EtOH |
93-97 |


a) The reaction should be carried out with using 90% (v/v) concd H2SO4, under the same remaining reaction conditions.
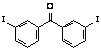
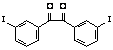
Finally, several slightly deactivated arenes, ArH, as well as benzil and diphenyl sulfone (Table 3) were ”directly” diiodinated at 25-30 oC for 120 minutes, with the iodinating solutions containing I+ intermediates in a 10% excess. After typical workups and purification, the purified diiodinated products were obtained in 17-72% yields. Only benzophenone should be diiodinated by the ”inverse” method at 0-5 oC for 120 minutes; otherwise, the crude diiodinated benzophenone was considerably admixed with various isomeric side products, difficult to remove from the desirable 3,3’-diiodobenzophenone; cf. Ref. 8, p. 9117.
Table 3. Diiodinated pure products prepared
|
ArH |
ArI |
Yield (%) |
Mp (oC) Solvent |
Lit, mp (oC) |
|
|
I2/NaIO4 |
KI/NaIO4 |
||||
|
|
|
71 |
72 |
333-334 EtOH |
334-335 |
|
|
|
32 |
33 |
264-266 EtOH |
255-256 |
|
|
|
62 |
62 |
317-318 EtOH |
not found |
|
|
|
71 |
68 |
289-290 EtOH |
303-304 288-290 |
|
|
|
40 |
40 |
301-302 EtOH |
289-290 300-302 |
|
|
|
17 |
19 |
135-139 EtOH |
not found |
|
|
|
55 |
57 |
117-118 EtOH |
115-116 |
|
|
|
33 |
32 |
131-132 EtOH |
133-135 |
|
|
|
31a |
32a |
150-152 Me2CO |
151-153 |
|
|
|
45 |
45 |
125-127 EtOH |
124-128 |
|
|
|
46 |
47 |
121-122 EtOH |
122-123 |





a) The reaction should be carried out with using the ”inverse” method at 0-5 oC and within 120 min.
By comparison with our former paper [8], the present communication interestingly extends our previous results. However, in order to further increase some low final yields reported in the Tables 1-3, it is necessary to carry out a number of additional reactions, with the iodinating solutions containing I+ intermediates in a 50% excess, whereas for nitrobenzene – in a 100% excess. The results thus extended will be published as soon as possible.
Experimental
General
The melting points of the freshly purified iodinated products are uncorrected (Table 1 – 3); they were microanalyzed (%I), and their 1H and 13C NMR spectra (not shown here) were recorded at r.t. with a Bruker AVANCE DMX 400 MHz spectrometer, in CDCl3 solutions, and they were next compared with the same spectra of authentic samples available in our laboratory [3 – 8]. All the reagents and solvents were commercial (Aldrich) and were used as such without further purification. Only elemental iodine (diiodine) should be finely powdered to facilitate its dissolution in the reaction mixtures.
Oxidative mono- or diiodination reactions of various arenes
Preparation of four iodinating solutions applied in the following reactions
I. Iodinating solutions used for the monoiodination of arenes (listed in the Tables 1 and 2)
a) with using I2/NaIO4/concd (min. 95%) H2SO4 system
Powdered diiodine (0.60 g; 2.36 mmol) and next NaIO4 (0.17 g; 0.79 mmol) were suspended in stirred concd (min. 95%) H2SO4 (15 mL). The suspension thus obtained was stirred at 25 – 30 °C for 30 min to afford a dark brown iodinating solution containing ca. 5.5 mmol of the I+ intermediates.
b) with using KI/NaIO4/concd (min. 95%) H2SO4 system
NaIO4 (0.29 g; 1.36 mmol) was dissolved in stirred concd (min. 95%) H2SO4 (15 mL). Next KI (0.68 g; 4.1 mmol) was slowly added portionwise, with stirring. The suspension thus obtained was stirred at 25 – 30 °C for 30 min to afford a dark brown iodinating solution containing ca. 5.5 mmol of the I+ intermediates.
II. Iodinating solutions used for the diiodination of arenes (listed in the Table 3)
c) with using I2/NaIO4/concd (min. 95%) H2SO4 system
Powdered diiodine (1.20 g; 4.72 mmol) and next NaIO4 (0.34 g; 1.59 mmol) were suspended in stirred concd (min. 95%) H2SO4 (30 mL). The suspension thus obtained was stirred at 25 – 30 °C for 30 min to afford a dark brown iodinating solution containing ca. 11 mmol of the I+ intermediates.
d) with using KI/NaIO4/concd (min. 95%) H2SO4 system
NaIO4 (0.58 g; 2.72 mmol) was dissolved in stirred concd (min. 95%) H2SO4 (30 mL). Next KI (1.36 g; 8.20 mmol) was slowly added portionwise, with stirring. The suspension thus obtained was stirred at 25 – 30 °C for 30 min to afford a dark brown iodinating solution containing ca. 11 mmol of the I+ intermediates.
“Direct” monoiodination of arenes listed in the Table 1
To the stirred iodinating solutions, either Ia) or Ib), containing ca 5.5 mmol of I+ intermediates (used in a 10% excess) individual arenes shown in the Table 1 (5 mmol; 0% excess) were added at once, and the mixtures were stirred at 25 – 30 °C for 60 min. The reactions were quenched by pouring the final reaction mixtures into stirred ice-water (150 g). The precipitates were collected by filtration, washed well with ice water until the filtrates were neutral, dried preliminarily by the suction, and next air-dried in the dark. To obtain the purified monoiodinated products, the solid crude products were recrystallized from appropriate organic solvents (Table 1).
The monoiodination of benzaldehyde should be carried out as above, but in 90% (v/v) conc. H2SO4. When the same reaction was carried out in concd (min. 95%) H2SO4, 3-IC6H4CHO was admixed by 3-IC6H4COOH.
“Inverse” monoiodination of arenes listed in the Table 2
Individual arenes shown in the Table 2 (5 mmol; 0% excess) were suspended, with stirring, in concd (min. 95%) H2SO4 (10 mL), then the stirred suspensions were cooled to 0 – 5 °C. Next iodinating solutions, either Ia) or Ib), containing ca 5.5 mmol of I+ intermediates (used in a 10% excess) were slowly added dropwise within 45 min, while keeping the same temperature. The stirring at 0 – 5 °C was continued for a further 15 min. The reactions were quenched by pouring the final reaction mixtures into stirred ice-water (200 g). The precipitates were collected by filtration, and the following workups and purifications were the same as above to obtain the purified monoiodinated products (Table 2).
The monoiodination of methyl anisoate should be carried out as above, but in 90% (v/v) conc. H2SO4. Otherwise, the monoiodinated crude product was notably contaminated and difficult for its purification.
“Direct” diiodination of arenes listed in the Table 3
To the stirred iodinating solutions, either IIa) or IIb), containing ca 11 mmol of I+ intermediates (used in a 10% excess) individual arenes shown in the Table 3 (5 mmol; 0% excess) were added at once, and the mixtures were stirred at 25 – 30 °C for 120 min. The reactions were quenched by pouring the final reaction mixtures into stirred ice-water (200 g). The precipitates were collected by filtration, and the following workups and purifications were the same as above to obtain the purified diiodinated products (Table 3).
The diiodination of benzophenone should be carried out by the “inverse” method at 0 – 5 °C. Otherwise, the diiodinated crude product was notably contaminated with isomeric side products; cf. Ref. 8, p. 9117, the right column.
References and Notes
1. a) Roedig, A. In Houben-Weyl, Methoden der Organischen Chemie, 1960, Vol. V/4, 517-678; b) Merkushev, E. B. Advances in the Synthesis of Iodoaromatic Compounds. Synthesis 1988, 923-937.
2. a) Varvoglis, A. The Organic Chemistry of Polycoordinated Iodine; VCH: Weinheim, 1992; b) Zhdankin, V. V.; Stang, P. J. Recent Development in the Chemistry of Polyvalent Iodine compounds. Chem. Rev. 2002, 102, 2523-2584; c) Hypervalent Iodine Chemistry, series Topics in Current Chemistry, Vol. 224; Wirth, T., Ed.; Springer: Berlin, 2003.
3. Skulski, L. Organic Iodine(I, III and V) Chemistry: 10 Years of Development at the Medical University of Warsaw, Poland (1990-2000). Molecules 2000, 5, 1331-1371. See pp 1332-1345. Avail at http://www.mdpi.org/molecules
4. Skulski, L. Novel Easy Preparations of Some Aromatic Iodine(I, III, and V) Reagents, Widely Applied in Modern Organic Synthesis. Molecules, 2003, 8, 45-52. Avail at http://www.mdpi.org/molecules
5. Krassowska-Swiebocka, B.; Lulinski, P.; Skulski, L. Chemical Manganese Dioxide (CMD): Its Application to Oxidative Iodination of Benzene, Halobenzenes and Some Deactivated Arenes. Molecules, 2004, 9, 595-601. Avail. at http://www.mdpi.org/molecules
6. Sosnowski, M., Skulski, L., Wolowik, K. Microwave-Accelerated or Conventionally Heated Iodination Reactions of Some Aromatic Amines, Using ortho-Periodic Acid as the Oxidant. Molecules 2004, 9, 617-621. Avail. at http://www.mdpi.org/molecules
7. Lulinski, P.; Kryska, A.; Sosnowski, M.; Skulski, L. Eco-friendly Oxidative Iodination of Various Arenes with a Urea-Hydrogen Peroxide Adduct (UHP) as the Oxidant. Synthesis 2004, 441-445.
8. Kraszkiewicz, L.; Sosnowski, M.; Skulski, L. Easy inexpensive and effective oxidative iodination of deactivated arenes in sulfuric acid. Tetrahedron 2004, 60, 9113-9119.
9. Dictionary of Organic Compounds, 6th ed.; Chapman & Hall: London, 1996.